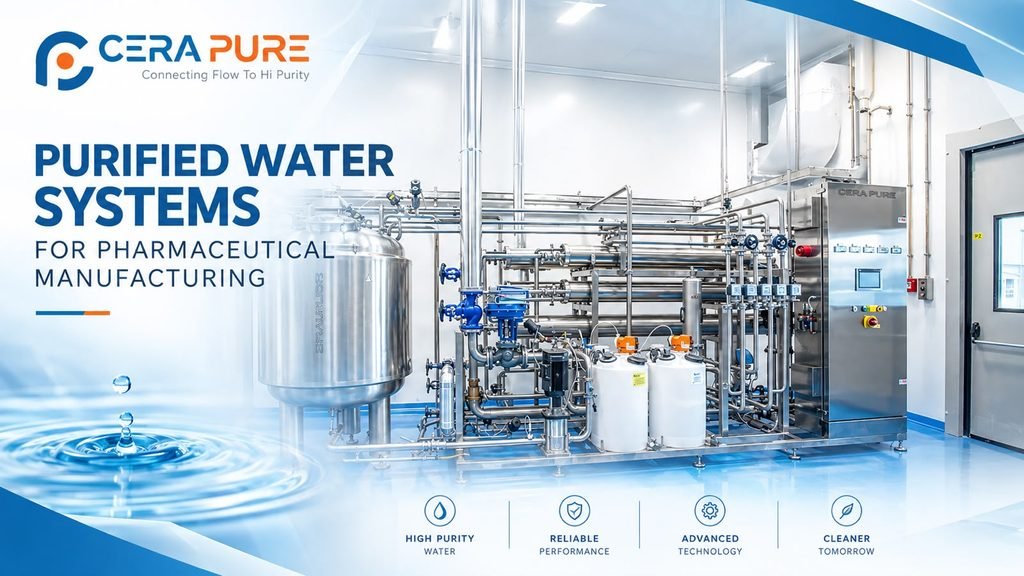
Clean and sterile manufacturing environments are essential in pharmaceutical and biopharmaceutical production. Process vessels, tanks, pipelines, transfer lines, filters and other product-contact equipment must be cleaned and, where required, sterilized before the next manufacturing operation.
Clean-in-Place (CIP) and Sterilization-in-Place (SIP) are widely used systems that allow pharmaceutical equipment to be cleaned and sterilized without dismantling the complete process system.
A properly designed CIP/SIP system can improve cleaning consistency, reduce manual intervention, minimize equipment downtime and support contamination-control requirements. However, simply installing a CIP or SIP skid is not enough. The complete process—including equipment design, flow, spray coverage, temperature, pressure, chemical concentration, drainability and validation—must work together.
What Are CIP and SIP in Pharma?
CIP – Clean-in-Place
CIP stands for Clean-in-Place.
It is a method of cleaning process equipment without dismantling it. Cleaning solutions, water and other fluids are circulated through tanks, vessels, pipelines and associated equipment according to a defined cleaning cycle.
The objective is to remove:
- Product residues
- Active pharmaceutical ingredients
- Excipients
- Oils and fats
- Proteins
- Cleaning-agent residues
- Microbial contamination
- Other process-related deposits
SIP – Sterilization-in-Place
SIP generally refers to Sterilization-in-Place and is commonly performed using clean/pure steam.
After suitable cleaning, steam is introduced into the process equipment and associated piping to achieve the required thermal sterilization conditions.
SIP is particularly important for sterile and aseptic manufacturing processes where product-contact equipment must achieve an appropriate level of sterility assurance.
CIP vs SIP: What Is the Difference?
| Parameter | CIP | SIP |
|---|---|---|
| Full Form | Clean-in-Place | Sterilization-in-Place |
| Main Purpose | Remove residues and contaminants | Reduce/eliminate viable microorganisms through validated sterilization |
| Typical Medium | Water, cleaning chemicals, rinse water | Clean/pure steam |
| Main Control Parameters | Flow, temperature, time, concentration | Temperature, pressure, time, steam quality |
| Common Applications | Tanks, vessels, pipelines, process equipment | Sterile vessels, process lines, filters and product-contact systems |
| Performed | Before SIP when both are required | Usually after cleaning |
| Validation Focus | Residue removal and cleaning effectiveness | Sterilization effectiveness and heat distribution |
Why Are CIP and SIP Important in Pharmaceutical Manufacturing?
Pharmaceutical manufacturing requires consistent control of contamination and cross-contamination risks.
Manual cleaning can introduce variability because cleaning effectiveness may depend on operator technique, access to difficult areas and the consistency of the cleaning procedure.
A properly engineered CIP/SIP system can provide:
- Repeatable cleaning cycles
- Reduced manual intervention
- Lower equipment dismantling requirements
- Reduced cleaning-related downtime
- Better process control
- Improved documentation
- Easier monitoring of critical parameters
- Better support for cleaning and sterilization validation
How Does a Pharmaceutical CIP System Work?
A typical CIP cycle may contain several stages. The exact sequence depends on the equipment, residue characteristics and validated cleaning procedure.
1. Pre-Rinse
The equipment is initially rinsed with water to remove loose and water-soluble residues.
2. Caustic or Detergent Wash
A suitable cleaning solution is circulated through the equipment. For example, alkaline cleaning agents may be used for certain organic or protein-based residues.
3. Intermediate Rinse
After chemical cleaning, the system is rinsed to remove loosened contaminants and residual cleaning chemicals.
4. Acid Cleaning
Where required, an acidic cleaning stage may be incorporated to remove mineral deposits, scale or other residues that are not effectively removed during alkaline cleaning.
5. Final Rinse
The system is thoroughly rinsed to remove remaining cleaning-agent residues. Depending on the process, purified water or WFI may be used for the final rinse.
6. Drainage and Drying
The equipment should be designed to drain effectively after cleaning. Retained water can create conditions for microbial growth, making drainability an important part of hygienic equipment design.
The Four Important Factors of an Effective CIP Cycle
Cleaning effectiveness is not determined by chemical concentration alone.
Time
The cleaning solution must remain in contact with the contaminated surface for sufficient time.
Temperature
Temperature can influence chemical reaction rates and the removal of certain residues. However, higher temperature is not automatically better because the process must remain compatible with the product residue, materials and equipment.
Chemical Concentration
The concentration of the selected cleaning agent must remain within the validated operating range.
Mechanical Action
Flow velocity, turbulence, spray coverage and hydraulic conditions contribute to the physical removal of residues.
CIP System Components
A pharmaceutical CIP system may include:
- CIP storage tanks
- CIP pumps
- Heat exchangers
- Spray balls and rotary spray devices
- Automated sanitary valves
- Temperature sensors
- Pressure transmitters
- Flow meters
- Conductivity sensors
- Level sensors
- pH measurement
- PLC/SCADA control systems
Why Are Dead Legs a Problem in CIP and SIP Systems?
A dead leg is an area of process piping or equipment where fluid or steam cannot circulate effectively.
Dead legs can create areas where:
- Product residues remain
- Cleaning chemicals do not reach adequately
- Rinse water becomes trapped
- Microorganisms can survive
- Steam penetration may be inadequate
- Drainage becomes difficult
Therefore, hygienic equipment design should minimize unnecessary dead legs and provide appropriate drainage and access.
Spray Coverage and CIP Cleaning
One important question in CIP design is whether the cleaning solution actually reaches every required product-contact surface.
A vessel may have a correctly sized CIP pump and still experience cleaning failures if the spray device does not provide adequate coverage.
Spray coverage can depend on:
- Vessel geometry
- Spray-ball design
- Spray-device location
- Flow rate
- Pressure
- Surface condition
- Internal components
- Baffles and agitators
- Nozzle arrangement
What Is CIP Cleaning Validation?
Cleaning validation is the documented evidence that a defined cleaning process consistently removes residues to predetermined acceptable levels.
A cleaning validation program may consider:
- Product residue
- Cleaning-agent residue
- Microbial contamination
- Endotoxin, where applicable
- Equipment configuration
- Worst-case products
- Worst-case locations
- Sampling methods
- Acceptance criteria
- Cleaning cycle parameters
How Is CIP Effectiveness Verified?
Visual Inspection
Visible residues or contamination can sometimes be identified through visual inspection.
Swab Sampling
Samples are collected from defined product-contact locations and analyzed for residues.
Rinse Sampling
The final rinse can be sampled and analyzed to determine whether residues remain within acceptable limits.
TOC Testing
Total Organic Carbon (TOC) can be used as an analytical indicator for certain organic residues.
Conductivity
Conductivity can help monitor cleaning-agent or ionic residue during rinse stages.
Microbiological Testing
Microbial testing may be relevant depending on the manufacturing process and contamination-control requirements.
What Is SIP in Pharmaceutical Manufacturing?
SIP, or Sterilization-in-Place, uses steam to sterilize equipment without dismantling the process system.
It is commonly used for:
- Pharmaceutical process vessels
- Biopharmaceutical equipment
- Product transfer lines
- Sterile process piping
- Filling-related equipment
- Filters and associated systems
- Other suitable product-contact equipment
SIP is generally performed after the equipment has been adequately cleaned because sterilization is not intended to replace effective cleaning.
How Does the SIP Process Work?
1. Heat-Up
Steam enters the system and gradually raises the temperature of the equipment and connected piping. Air removal and effective steam distribution are critical because trapped air can interfere with steam contact.
2. Sterilization Hold
Once the required conditions are achieved at the defined critical locations, the system is maintained for the validated exposure period.
3. Cooling
After sterilization, the equipment is cooled under controlled conditions. The sterile boundary should be maintained during cooling to minimize the possibility of recontamination.
Important SIP Design Considerations
An effective SIP system should consider:
- Steam distribution
- Air removal
- Condensate removal
- Drainability
- Temperature distribution
- Pressure control
- Steam quality
- Instrument calibration
- Sterile boundary protection
- Equipment integrity
- Filter integrity, where applicable
- Automation and data recording
How Is SIP Validated?
SIP validation should demonstrate that the required sterilization conditions are achieved at the locations that are hardest to sterilize.
Validation can involve:
- Temperature mapping
- Heat distribution studies
- Identification of cold spots
- Cycle development
- Biological indicators, where appropriate
- Sterilization lethality calculations
- Steam quality evaluation
- Instrument calibration
- Repeatability studies
Role of WFI and Purified Water in CIP
Water quality is an important consideration in pharmaceutical cleaning processes.
Depending on the application, Purified Water (PW) or Water for Injection (WFI) may be used at different stages.
The selection depends on:
- Product requirements
- Manufacturing process
- Equipment design
- Cleaning procedure
- Regulatory requirements
- Final rinse requirements
CIP and SIP Automation
Modern pharmaceutical CIP/SIP systems can be integrated with PLC and SCADA platforms.
Automation can help control and record:
- Flow rate
- Temperature
- Pressure
- Chemical concentration
- Conductivity
- Cycle duration
- Valve sequence
- Tank level
- Alarm conditions
- Recipe selection
Common CIP Problems in Pharmaceutical Plants
Inadequate Spray Coverage
The cleaning solution does not adequately contact all internal surfaces.
Insufficient Flow
The required mechanical cleaning action may not be achieved.
Incorrect Chemical Concentration
A cleaning agent outside its validated concentration range can reduce cleaning effectiveness.
Poor Temperature Control
Incorrect temperature can affect cleaning performance.
Dead Legs
Poor piping design can create areas that are difficult to clean or drain.
Poor Drainability
Retained cleaning solution or water can create contamination risks.
Incorrect Valve Routing
An incorrect automated valve sequence can prevent cleaning solution from reaching the intended circuit.
Inadequate Documentation
A technically effective process can still create qualification or audit challenges when critical operating and validation documentation is incomplete.
CIP/SIP System Design Checklist
| Design Area | Key Consideration |
|---|---|
| Equipment Geometry | Minimize difficult-to-clean areas |
| Piping | Hygienic routing and appropriate slope |
| Dead Legs | Minimize and assess critical locations |
| Spray Devices | Ensure suitable surface coverage |
| Flow | Maintain required cleaning flow |
| Temperature | Control and record critical temperatures |
| Chemicals | Define validated concentration ranges |
| Drainability | Prevent retained water and chemicals |
| Steam | Ensure appropriate steam distribution |
| Instrumentation | Calibrated critical sensors |
| Automation | Controlled and repeatable recipes |
| Validation | Document cleaning and sterilization studies |
| Documentation | Maintain drawings, protocols and records |
CIP vs Manual Cleaning
| Factor | Manual Cleaning | CIP |
|---|---|---|
| Manual Intervention | Higher | Lower |
| Repeatability | Operator dependent | Recipe controlled |
| Equipment Dismantling | Often required | Usually minimized |
| Documentation | More manual | Can be automatically recorded |
| Cleaning Consistency | Can vary | More repeatable when validated |
| Large Fixed Systems | Difficult | More suitable |
| Validation | Can be more difficult to standardize | Can be validated around defined cycles |
When Should CIP and SIP Be Used Together?
CIP and SIP are often integrated where a manufacturing process requires both cleaning and sterilization.
A simplified sequence can be:
Production → Pre-Rinse → Chemical Cleaning → Intermediate Rinse → Final Rinse → Drain/Dry → SIP → Controlled Cooling → Production
The actual sequence should be established based on the equipment, process, product and validated procedures.
Benefits of a Properly Designed CIP/SIP System
A well-designed and validated system can provide:
- Consistent cleaning performance
- Improved contamination control
- Reduced manual intervention
- Reduced equipment dismantling
- Repeatable process cycles
- Better process monitoring
- Reduced cleaning-related downtime
- Improved documentation
- Better support for GMP compliance
- Improved equipment utilization
CIP/SIP and GMP Compliance
Important elements include:
- Hygienic equipment design
- Documented cleaning procedures
- Defined operating parameters
- Appropriate validation protocols
- Calibrated instruments
- Cleaning verification
- Sterilization validation where applicable
- Proper documentation and data recording
- Periodic review of validated processes
- Change control and requalification when required
Frequently Asked Questions About CIP and SIP
What is CIP in pharma?
CIP means Clean-in-Place. It is a method of cleaning pharmaceutical process equipment without dismantling the equipment, using controlled circulation of water and cleaning solutions.
What is SIP in pharma?
SIP means Sterilization-in-Place. It generally uses steam to sterilize pharmaceutical process equipment without dismantling the system.
What is the difference between CIP and SIP?
CIP primarily removes product residues and contaminants, while SIP is used to achieve validated sterilization conditions for suitable equipment and processes.
Is CIP required before SIP?
When both processes are used, cleaning is normally performed before sterilization because SIP does not replace effective removal of product residues.
How is CIP validated?
CIP validation can include documented cycle parameters, visual inspection, swab sampling, rinse sampling and analytical testing such as TOC or conductivity, depending on the process.
How is SIP validated?
SIP validation can involve temperature mapping, heat distribution studies, cold-spot assessment, sterilization-cycle development and other appropriate validation methods.
Why are dead legs important in CIP/SIP?
Dead legs can prevent effective circulation, cleaning, steam penetration and drainage. Poorly designed dead legs can therefore become difficult-to-clean or difficult-to-sterilize locations.
What water is used for pharmaceutical CIP?
Purified Water or WFI may be used depending on the cleaning stage, equipment and process requirements.
Does every pharmaceutical process require SIP?
No. SIP requirements depend on whether the manufacturing process requires in-place sterilization and the applicable process and contamination-control requirements.
Conclusion
CIP and SIP are important technologies for pharmaceutical and biopharmaceutical manufacturing where controlled cleaning and sterilization of process equipment are required.
CIP focuses on effective and repeatable removal of residues, while SIP focuses on validated sterilization of suitable equipment. Their performance depends not only on the CIP/SIP skid but also on equipment geometry, piping design, spray coverage, flow, temperature, chemical concentration, steam distribution, drainage, instrumentation and validation.
For pharmaceutical facilities, the most effective approach is to design CIP and SIP as part of the complete process system rather than treating them as standalone utilities. Proper engineering, automation, documentation and validation help establish repeatable cleaning and sterilization performance.